
Webinar
The integrated stress response promotes immune evasion through lipocalin 2

0:00
Abstract
Cancer cells activate the integrated stress response (ISR) to adapt to stress and resist therapy. ISR signals converge on activating transcription factor 4 (ATF4), which controls cell-intrinsic transcriptional programs that are involved in metabolic adaptation, survival and growth. However, whether the ISR–ATF4 axis influences anti-tumour immune responses remains mostly unknown. Here we show that loss of ATF4 decreases tumour progression considerably in immunocompetent mice, but not in immunocompromised ones, by enhancing T cell-dependent anti-cancer immune responses. An unbiased genetic screen of ATF4-regulated genes identifies lipocalin 2 (LCN2) as the principal ATF4-dependent effector that impairs anti-tumour immunity by favouring infiltration with immunosuppressive interstitial macrophages. Furthermore, we find that LCN2 promotes T cell exclusion and immune evasion in preclinical mouse models, and correlates with decreased T cell infiltration in patients with lung and pancreatic adenocarcinomas. Anti-LCN2 antibodies promote robust anti-tumour T cell responses in mouse models of aggressive solid tumours. Our study shows that the ATF4–LCN2 axis has a cell-extrinsic role in suppressing anti-cancer immunity, and could pave the way for an immunotherapy approach that targets LCN2.
What you'll learn
- How cancer cells can reshape the tumor immune environment and suppress anti-tumor responses.
- How a newly discovered pathway connecting macrophages and T cell exclusion in immune evasion, providing new insight into interactions between tumors and immune cells.
- How cell avidity can help uncover biologically relevant interactions, including the use of primary macrophages and a protein-on-bead approach to investigate cellular binding.
Technical note:
Get 5 of each and filter based on business unit client side.
This shows 5 of each card, and then filters them client side on BU.
Webinar, Scientific update, Whitepaper, Application note, Brochure.
We only show 4 cards in the end.
In design only 1 is shown, but the rest will be loaded when published.
Get 5 of each and filter based on business unit client side.
This shows 5 of each card, and then filters them client side on BU.
Webinar, Scientific update, Whitepaper, Application note, Brochure.
We only show 4 cards in the end.
In design only 1 is shown, but the rest will be loaded when published.
Thank you! Your submission has been received!
Oops! Something went wrong while submitting the form.
Button Text
The integrated stress response promotes immune evasion through lipocalin 2
Webinar
November 3, 2026
CA
Cancer cells activate the integrated stress response (ISR) to adapt to stress and resist therapy. ISR signals converge on activating transcription factor 4 (ATF4), which controls cell-intrinsic transcriptional programs that are involved in metabolic adaptation, survival and growth. However, whether the ISR–ATF4 axis influences anti-tumour immune responses remains mostly unknown. Here we show that loss of ATF4 decreases tumour progression considerably in immunocompetent mice, but not in immunocompromised ones, by enhancing T cell-dependent anti-cancer immune responses. An unbiased genetic screen of ATF4-regulated genes identifies lipocalin 2 (LCN2) as the principal ATF4-dependent effector that impairs anti-tumour immunity by favouring infiltration with immunosuppressive interstitial macrophages. Furthermore, we find that LCN2 promotes T cell exclusion and immune evasion in preclinical mouse models, and correlates with decreased T cell infiltration in patients with lung and pancreatic adenocarcinomas. Anti-LCN2 antibodies promote robust anti-tumour T cell responses in mouse models of aggressive solid tumours. Our study shows that the ATF4–LCN2 axis has a cell-extrinsic role in suppressing anti-cancer immunity, and could pave the way for an immunotherapy approach that targets LCN2.
This is some text inside of a div block.

The CAR Immune Synapse Interactome: How Membrane Organization Shapes CAR T–Tumor Cell Interactions
Webinar
September 14, 2026
CA
Unlike conventional drugs, whose activity is shaped by binding to a single target, CAR T-cell (CART) engagement with tumor cells involves multiple receptors, adhesion molecules, and signaling proteins. These components are partially organized within cholesterol-rich membrane microdomains called lipid rafts, forming the CAR immune synapse (CARIS)—the interface through which a CART recognizes, attaches to, and kills a tumor cell.
To evaluate CARIS as a pharmacological interface, we compared CD28- and 4-1BB-CARTs, which exhibit distinct therapeutic profiles. The rapid antitumor activity of CD28-CARTs was associated with brief, dynamic synapses supporting rapid signaling and serial killing. Conversely, 4-1BB-CARTs formed longer, mechanically stable synapses consistent with their slower but sustained activity. Synapse strength was measured using z-Movi acoustic force microscopy.
Lipid rafts were enriched at CARIS. Attenuating membrane lipid raft cholesterol preferentially compromised 4-1BB CARIS mechanical stability, while predominantly impairing CD28 CARIS activation and cytolytic output. The molecular networks underlying these differences were mapped by integrating lipid-raft lipidomics, differential SILAC proteomics to distinguish CART-derived from tumor-derived components, and T-cell transcriptomics.
CD28-associated lipid rafts were enriched in diacylglycerol signaling components, PKCθ, small GTPases, Src-family kinases, and AKT/PI3K-proximal modules, supporting rapid signal-to-function coupling. Stomatin-family and flotillin scaffolds favored dynamic remodeling, while CD8-dominant transcriptional programs supported cytolytic degranulation. Engaged tumor cells displayed membrane-stress signatures consistent with late-stage senescence and apoptotic commitment.
In contrast, 4-1BB-associated lipid rafts were enriched in lysophosphatidic acid, an inhibitory lipid known to suppress CD8 effector functions while promoting integrin-dependent adhesion, consistent with the marked enrichment of integrins at CARIS, NF-κB and cytokine signaling components, and ERM scaffolds that couple lipid rafts to cortical actin, sustaining CARIS stability. Engaged tumor cells exhibited signatures of early apoptotic resistance, including damage-response and repair markers.
Together, these findings establish CARIS as a central determinant of CART function and therapeutic behavior. Mapping and manipulating its lipid–protein networks provide a platform for understanding CART pharmacodynamics and developing strategies to improve therapeutic efficacy.
This is some text inside of a div block.


Avidity and functional characterization of CD19/BCMA dual-targeting CAR-T cell designs in NHL
Webinar
July 1, 2026
CA
CD19-directed CAR-T therapy has achieved remarkable clinical outcomes in patients with relapsed or refractory non-Hodgkin lymphoma (NHL). Nevertheless, a significant proportion of patients develop primary resistance or relapse, often associated with CD19 antigen loss or downregulation. Given the co-expression of CD19 and BCMA in NHL, we hypothesized that dual antigen targeting could improve therapeutic durability and mitigate antigen escape. To address this, we developed a panel of CD19/BCMA dual-targeting CAR-T cells building on our academic platforms targeting CD19 (varnimcabtagene autoleucel, ARI0001) and BCMA (cesnicabtagene autoleucel, ARI0002h). Multiple strategies were explored, including pooled mono-targeted products, co-transduction with two lentiviral vectors, bicistronic constructs, and tandem/loop CAR designs incorporating dual binding domains within a single receptor. Functional activity and avidity were assessed across varying antigen expression conditions. We found that dual CAR-T cells generated through co-transduction consistently showed superior performance relative to single-target CD19 CAR-T cells and other dual-targeting formats, particularly in models with low CD19 expression. A first-in-human phase I clinical trial (CARTDBG-01; NCT06097455) is currently underway to evaluate the safety and efficacy of ARI0003 in NHL.
Related publication: Bachiller, M., Barceló-Genestar, N., Rodriguez-Garcia, A., Alserawan, L., Dobaño-López, C., Giménez-Alejandre, M., ... & Guedan, S. (2025). ARI0003: Co-transduced CD19/BCMA dual-targeting CAR-T cells for the treatment of non-Hodgkin lymphoma. Molecular Therapy, 33(1), 317-335. https://doi.org/10.1016/j.ymthe.2024.11.028
This is some text inside of a div block.
Revolutionize binding for the future of immunotherapy: A virtual Avidion demo
Webinar
April 15, 2026
CA
90% of immuno-oncology drugs entering Phase I clinical trials fail, due to a poor predictive value of current preclinical assays. To solve this, scientists and drug developers are turning to Cell Avidity, the total binding strength between cells, to better predict therapeutic efficacy.
Join us for an exclusive virtual demonstration of Avidion, the next generation of Cell Avidity technology. This session isdesigned for researchers and groups who understand the "why" of avidity but want to see the "how."
We will take you inside the instrument to explore the automated workflow, preview our intuitive software interface, and show real-world customer proof that validates its predictive power. See how Avidion transforms complexcellular interactions into actionable insights with push-button simplicity.
We know your time is valuable, that's why we designed this 30-minute intensive session specifically for researchers who need to seethe technology in action. Don’t miss your chance to talk directly with LUMICKS scientists and product managers. See the future of cell interaction analysis from your desk.
This is some text inside of a div block.

Enhancing efficacy against clear cell renal cell carcinoma through format-tuning of bispecific T cell engagers
Scientific update
January 29, 2025
CA
This is some text inside of a div block.
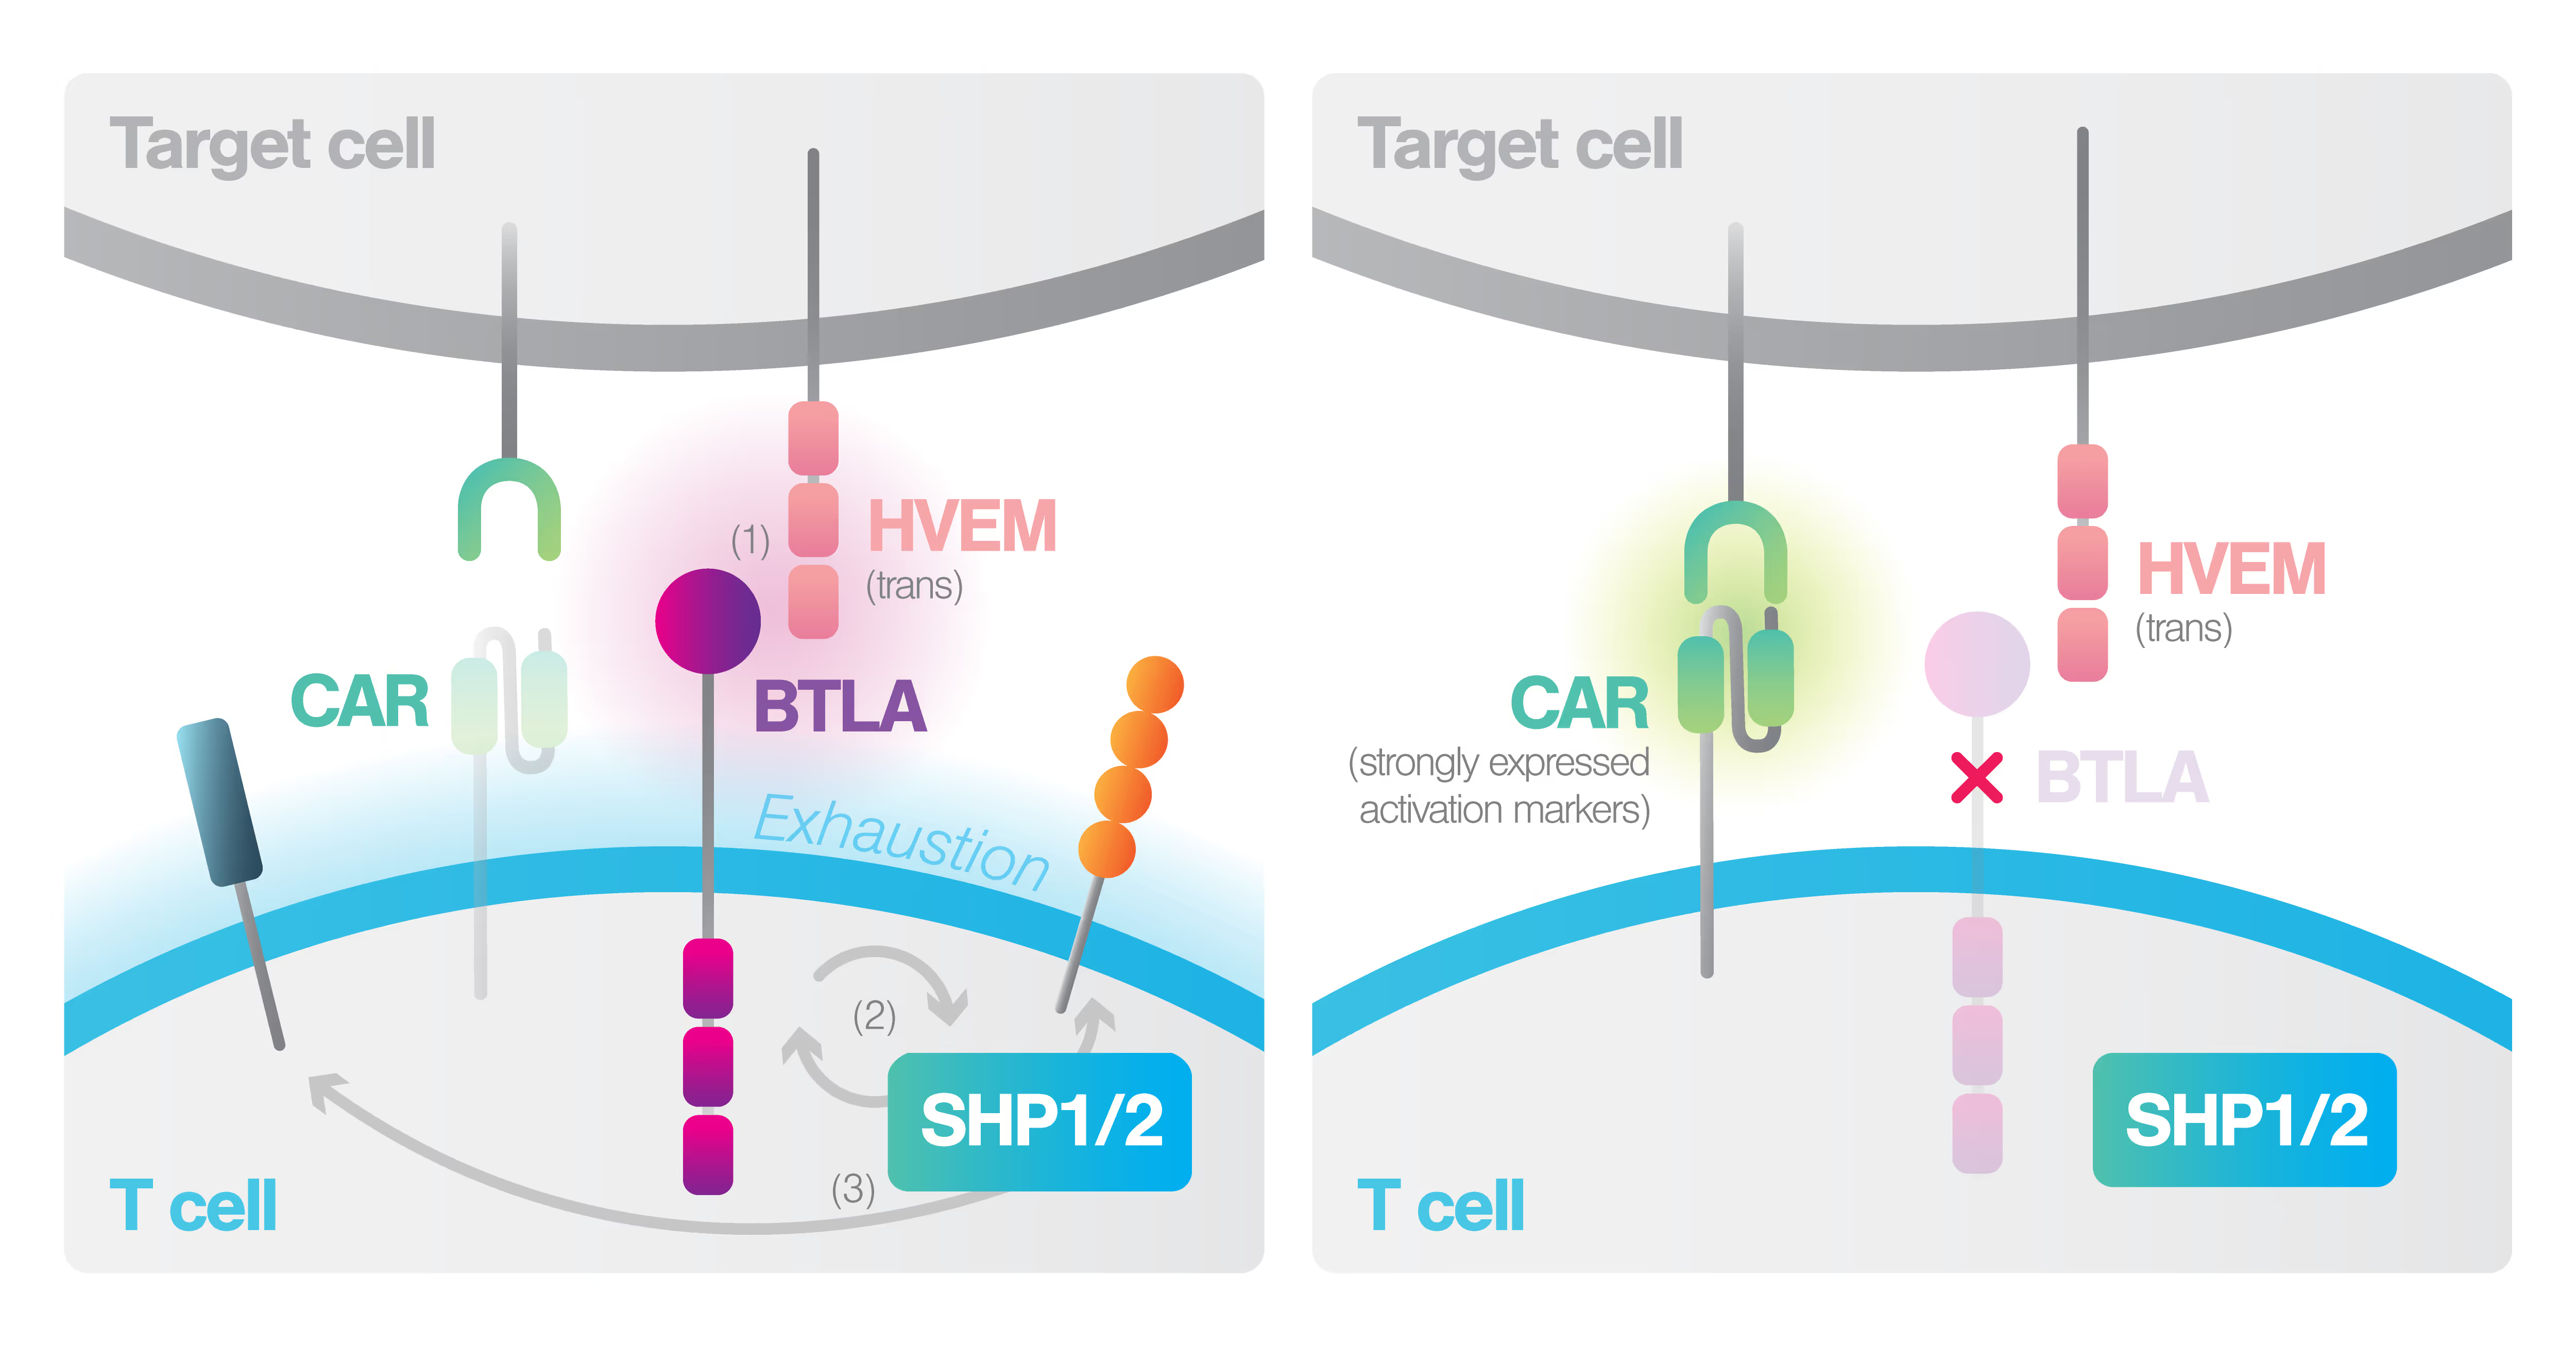
Cell Avidity uncovers critical binding mechanism impairing T-cell potency in the Tumor Microenvironment
Scientific update
August 1, 2024
CA
This is some text inside of a div block.
Cell Avidity screening deployed by T cell pioneers to accelerate the identification of functional TCRs
Scientific update
June 25, 2024
CA
This is some text inside of a div block.
Cell Avidity-tuning ensures safe CAR-T therapy for solid tumors
Scientific update
June 5, 2024
CA
This is some text inside of a div block.
Cell Avidity: a key to accelerate IND filing in cell therapy drug development
Whitepaper
July 1, 2023
CA
This is some text inside of a div block.
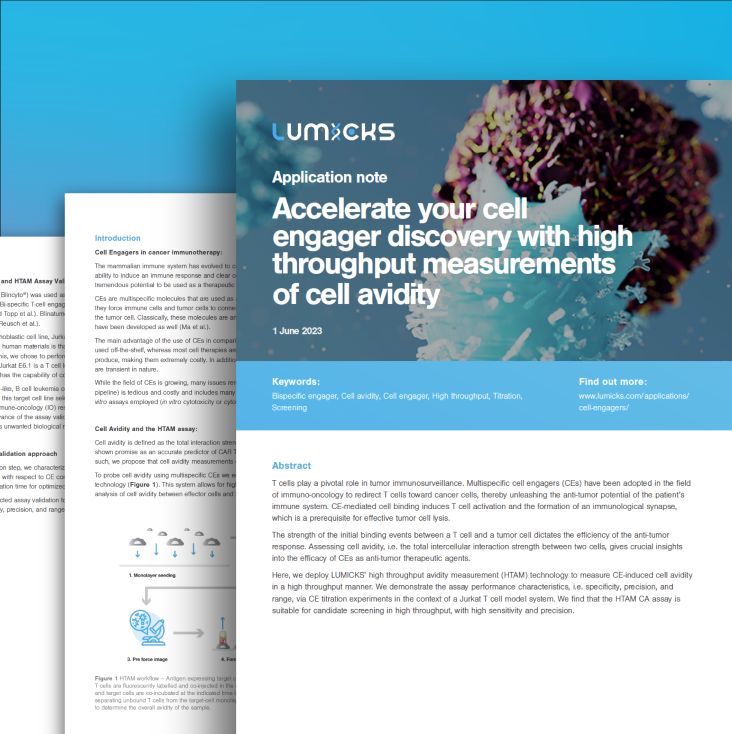
Accelerate your cell engager discovery with high throughput measuremenets of Cell Avidity
Whitepaper
June 1, 2023
CA
This is some text inside of a div block.

Accelerate your cell engager discovery with high throughput measurements of Cell Avidity
Application note
June 1, 2023
CA
T cells play a pivotal role in tumor immunosurveillance. Multispecific cell engagers (CEs) have been adopted in the field of immuno-oncology to redirect T cells toward cancer cells, thereby unleashing the anti-tumor potential of the patient’s immune system. CE-mediated cell binding induces T cell activation and the formation of an immunological synapse, which is a prerequisite for effective tumor cell lysis.
The strength of the initial binding events between a T cell and a tumor cell dictates the efficiency of the anti-tumor response. Assessing cell avidity, i.e. the total intercellular interaction strength between two cells, gives crucial insights into the efficacy of CEs as anti-tumor therapeutic agents.
Here, we deploy LUMICKS’ high throughput avidity measurement (HTAM) technology to measure CE-induced cell avidity in a high throughput manner. We demonstrate the assay performance characteristics, i.e. specificity, precision, and range, via CE titration experiments in the context of a Jurkat T cell model system. We find that the HTAM CA assay is suitable for candidate screening in high throughput, with high sensitivity and precision.
This is some text inside of a div block.
Identify functionally optimal TCR T cells through Cell Avidity measurements
Application note
June 7, 2021
CA
T cell receptor (TCR) -based cancer immunotherapy has the potential to become a powerful approach to treat solid tumors, such as melanoma. However, conventional methods that validate the effectiveness of TCR transduced T cells are often inconsistent with functional assays or are tedious to perform. In this application note, we show how the z-Movi® Cell Avidity Analyzer reliably and quickly identifies functionally optimal TCR-engineered T cells targeting melanoma cell lines based on cell–cell interaction strength (cellular avidity).
This is some text inside of a div block.
Rapid assessment of CAR T cell strategies for multiple myeloma with the z-Movi Cell Avidity Analyzer
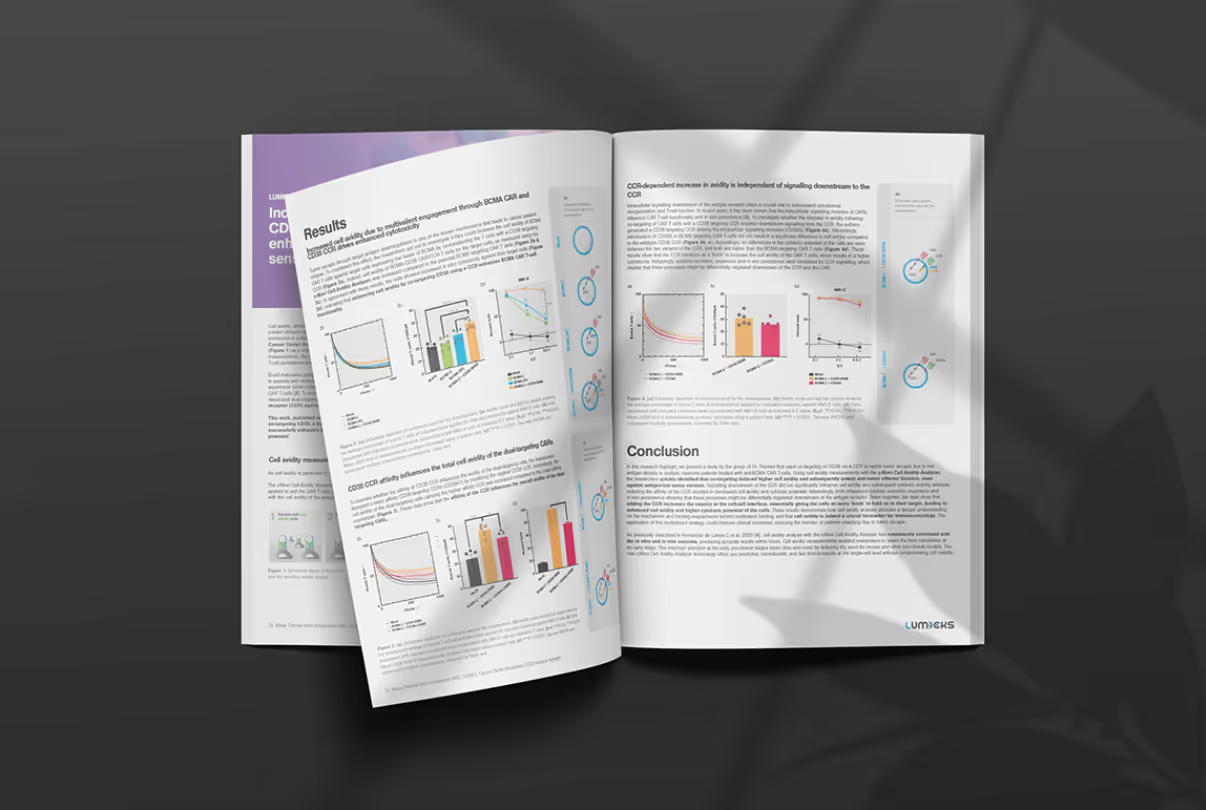
Rapid assessment of CAR T cell strategies for multiple myeloma with the z-Movi Cell Avidity Analyzer
Application note
June 5, 2020
CA
Identifying effective immunotherapeutic treatment strategies for multiple myeloma that also mitigate relapse often requires tedious and time-consuming validations, such as cell-killing assays and in vivo engraftments. We show that intercellular binding strength (cell avidity), measured by the z-Movi® Cell Avidity Analyzer, quickly predicts CAR T-cell efficacies that correlate with treatment outcomes in vivo.
This is some text inside of a div block.